

 menu


metal-organic compounds
 | IUCrDATA |
 access
4-Bromopyridinium trichloridostannate(II)

aChemistry, Osnabrück University, Barabarstr. 7, 49069 Osnabrück, Germany
*Correspondence e-mail: [email protected]
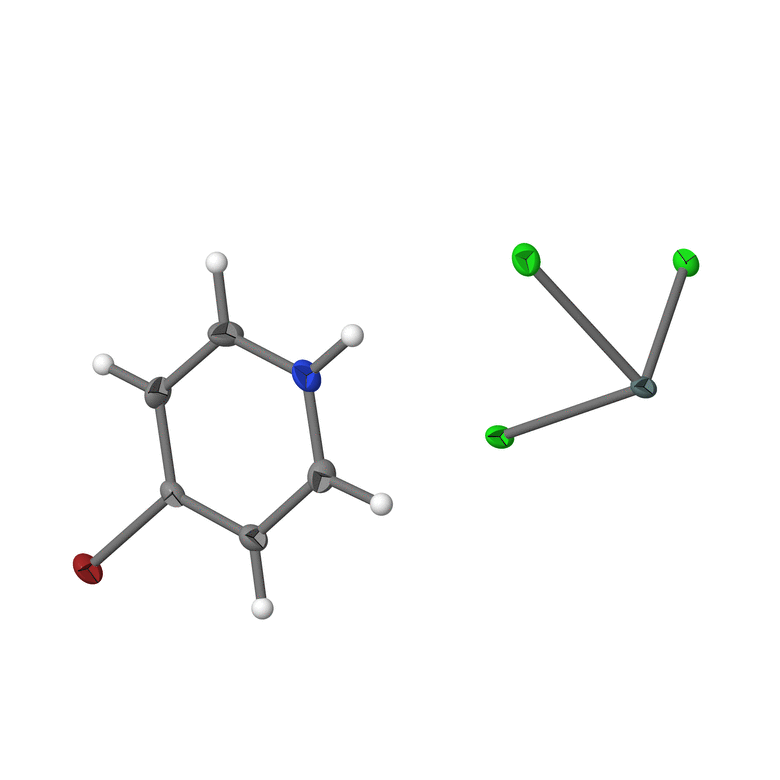
Single crystals of the title salt, (C5H5BrN)[SnCl3] or [4-BrPyH][SnCl3], were prepared from aqueous solutions of 4-bromopyridinium hydrochloride, [4-BrPyH]Cl, and tin dichloride, SnCl2. The cation has bond lengths and angles similar to those found in its other salts. In its first coordination sphere, the anion has a trigonal–pyramidal, tpy, structure with the SnII atom at the apex and the three chlorine atoms at the base. The tin–chlorine bond lengths are all greater than 2.5 Å and the bond angles are all less than 92°, which indicates the presence of tetrel bonds. These occur along the twofold screw-axis in the b-axis direction and extend the coordination sphere of the tin atom to a highly distorted, 33-aaa octahedron. In the crystal, the cations are stacked parallel to one another at varying distances along the a-axis direction, with weak π–π interactions occurring on both sides. Additional N—H⋯Cl hydrogen bonds are bifurcated and weak.
Keywords: crystal structure; trichloridostannate(II)-ion; 4-bromopyridinium; tetrel bonds; π–π interaction.
CCDC reference: 2582901
Structure description
The 4-bromopyrinium ion, [4-BrPyH]+, is not only a cation frequently used in connection with transition-metal anions such as [CuCl4]2− (Willett et al., 2003 ), [RuCl6]2− (Guthrie et al., 2026), [FeCl4]− (Zordan et al., 2005) and [CoHal4]− (Minguez Espallargas et al., 2006), but also when it comes to crystallizing anions of low-valent p-block elements. Typical examples in this case are anions of trivalent antimony, such as [SbIII2Hal9]3− (Nicolas et al., 2022), trivalent bismuth, such as [BiIII2Hal10]4− (Adonin et al., 2018) or tetravalent tellurium, such as [TeIVI6]2− (Walusiak et al., 2023). In this series of compounds, the title compound 4-bromopyridinium trichloridostannate(II), [4BrpyH][SnIICl3], is the first to contain an anion of divalent tin.
The asymmetric unit (Fig. 1 and Table 1) comprises one formula unit of each ion with all atoms in general positions. The cation exhibits the well-known structural features evident from the preceding publications: (i) the backbone of the pyridinium ion is nearly planar [Δ = ±0.011 (5) Å], (ii) the bromine atom lies slightly outside [0.0921 (5) Å] this plane, (iii) the bond angle at the nitrogen atom is significantly widened [122.7 (4)°], (iv) the bond angles at the carbon atoms in the meta position relative to the nitrogen atom are significantly narrowed [117.8 (5)/118.5 (4)°], (v) within the ring, the N—C bond lengths [1.330 (7)/1.340 (6) Å] are shorter than the C—C bonds [mean value: 1.383 (6) Å] and (vi) a Br—C distance of 1.873 (5) Å.
| ||||||||||||||||||||||||||
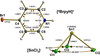 | Figure 1 The [4-BrPyH]+ and [SnCl3]− ion of the with atom numbering, bond lengths (Å), bond angles (°) and distance of the tin atom from the basal plane in square brackets. With the exception of the hydrogen atoms, which are shown as spheres of arbitrary radius, all other atoms are drawn as anisotropic displacement ellipsoids at the 50% probability level. |
With respect to the first coordination sphere, in which the SnII atom (Fig. 1) achieves an electron octet, the [SnCl3]− ion exhibits a trigonal–pyramidal, tpy, structure, with the tin atom at the apex and the three chlorine atoms in the basal plane. Given bond angles of around 90°, it can be assumed that the three empty, orthogonal 5p orbitals of the tin(II) atom are involved in the bonds.
Based on their structural parameters, the trichloridostannate(II) ions ions can be divided into two groups: in the first group, the tin–chlorine bond lengths are with a few exceptions shorter than 2.50 Å and the bond angles are all greater than 92°. This group includes, for example, the trichloridostannate(II) ions found in the compounds [Ph4P][SnCl3] (Müller et al., 1982), [SnIICl(18-crown-6)][SnCl3] (Drew & Nicholson, 1986) and [HfIV(OtBu)3(thf)3][SnCl3] (Njua et al., 2010). In the second group, one, two or all three tin–chlorine bonds are longer than 2.50 Å and one or more bond angles are less than 90°. Typical examples are: [MPyrH][SnCl3] (MPyrH = 1-methylpyrrolidin-1-ium; Liu et al., 2019), [Li(12-crown-4)(thf)][SnCl3] (Izod et al., 2012) and [tBuNH3][SnCl3] (Veith et al., 1988). In the first group, these are "naked" ions without strong intermolecular interactions involving the SnII atom with other building units; in the second group, they are ions in which the SnII atoms form additional so-called tetrel bonds (Bauzá et al., 2019; Brammer et al., 2023), characterized by the fact that they occur in a trans position to existing strong 2c–2e bonds. In the broadest sense, tetrel bonds can be regarded as extensive 3c–4e bonds based on an almost linear alignment of the p-orbitals of the three atoms involved in the X—Sn⋯Y arrangement (Reuter, 2025).
Based on the structural parameters observed in present case, the [SnCl3]− ion must, unambiguously, be classified in the second group. A look inside the crystal (Fig. 2) reveals that the anions are arranged in rows along the b axis with alternating orientations caused by a twofold crystallographic screw axis in the b-axis direction (Fig. 3), while the cations are stacked in a staggered arrangement along the a axis. In the case of the anions, the interactions result in three tetrel bonds (Fig. 3a, Table 2). With an asymmetry quotient Q = d(Sn⋯Cl)/d(Sn—Cl) = 1.365 (Cl3), 1.294 (Cl2) and 1.233 (Cl1), these 3c–4e bonds are highly asymmetric, causing the tpy, 30 configuration (Schröder et al., 2024), of the first coordination sphere being extended to a highly distorted octahedral, 33-aaa configuration. Similarly, the inter-penetration indices ρ (Echeverriá & Alvarez, 2023) using the van der Waals radii [rvdW(Cl) = 1.75 Å and rvdW(Sn) = 2.17 Å; Cordero et al., 2008] and the covalent radii [rcov(Cl) = 1.02 Å and rcov(Sn) = 1.39 Å; Mantina et al., 2009] support this concept. Furthermore, the calculation of bond valences, BS, according to Brese & O'Keeffe (1991) [Rij = 2.36, b = 0.37] shows that the intermolecular Sn⋯Cl distances contribute only slightly [0.23] to the calculated bond valence sum [BVScal = 1.96] of the divalent tin atom.
|
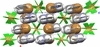 | Figure 2 Perspective view into the crystal of [4-BrPyH][SnCl3] looking down the b axis. All components are drawn as stick models with the cations in the background additionally represented as space-filling models. Colour code and van der Waals radii used: Sn = bronze, Cl = green, C = dark grey, 1.70 Å, H = white, 1.20 Å, N = light- blue, 1.55 Å, Br = brown, 1.85 Å. |
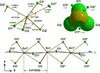 | Figure 3 Different representation of the tetrel bonds linking the [SnCl3]− ions. (above, left) ball-and-stick model of the tin environment with geometric parameters (Å, °) and asymmetry parameters Q (values in square brackets) characterizing the tetrel bonds, (above, right) space-filling model visualizing the inter-penetration of the van der Waals radii of tin and chlorine as result of the tetrel bond formation and interpenetration indices p (green values), (below) side-view on the twofold screw axis causing the tetrel-bond formation with the distance (Å) between two neighbouring tin atoms. Symmetry codes used to generate equivalent atoms: (1)  − x,  + y, − z; (2) − x, − + y, − z. |
The 4-bromopyridinium ions are stacked exactly parallel to one another, as they are flanked on both sides by crystallographic centres of symmetry at (0,0,0; Wyckhoff letter: a) and (1/2,0,0; Wyckhoff letter: d). The overlap of the pyridinium units, and thus the π–π interaction, is limited to the nitrogen atoms and two neighbouring carbon atoms (Fig. 4a). In this context, the extent of the overlap correlates with the distances between the least-squares planes defined by the ring atoms of neighbouring molecules (Fig. 4b): in the case of the centres of symmetry at a, the distance is significantly shorter (3.08 Å) and the overlap smaller than in the case of the centers of symmetry at d (3.84 Å) where the overlap is larger.
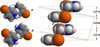 | Figure 4 Different representations of the cation stacking. (left) Space-filling model with stick model overlayed viewed perpendicular to the least-squares planes showing the extend of the overlap as resulting from the different centres of symmetry in Wyckoff positions a, and d. (right) Space-filling model of the stacking sequence in side-view with distances [Å] between the least-squares planes; from the two different centres of symmetry (black dots) responsible for the stacking sequence only the one at (1/2,1/2,1/2), a is visible. |
In addition to their contribution to the tetrel bonds, the chlorine atoms, Cl2 and Cl3, are involved into hydrogen bonds with the hydrogen atom attached to the nitrogen atom of the 4-bromopyridinium ion (Table 3). The donor–acceptor distances are quite long, as the hydrogen bond is bifurcated.
| ||||||||||||||||||||||
 .
Synthesis and crystallization
Stannous chloride (0.146 g (0.77 mmol); Sigma-Aldrich) was dissolved in approximately 10 ml of water together with 0.150 g (0.77 mmol) of 4-bromopyridine hydrochloride. Upon slow evaporation of the aqueous solutions at room temperature, the title compound crystallized out in the form of colourless blocks. Yield: 0.256 g (86% of the theoretical yield).
Refinement
Crystal data, data collection and structure details are summarized in Table 4. The maximum and minimum residual electron density peaks of 1.40 and −0.91 e Å−3, respectively, were located 1.99 and 1.97 Å from the H6 and Sn1 atoms, respectively.
|
Structural data
CCDC reference: 2582901
contains datablock I. DOI: https://doi.org/10.1107/S2414314626008771/tk4130sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S2414314626008771/tk4130Isup2.hkl
| (C5H5BrN)[SnCl3] | F(000) = 712 |
| Mr = 384.05 | Dx = 2.490 Mg m−3 |
| Monoclinic, P21/n | Mo Kα radiation, λ = 0.71073 Å |
| a = 7.2468 (2) Å | Cell parameters from 9904 reflections |
| b = 8.0253 (3) Å | θ = 2.3–27.5° |
| c = 17.7393 (6) Å | µ = 7.12 mm−1 |
| β = 96.834 (2)° | T = 100 K |
| V = 1024.35 (6) Å3 | Bloc, colourless |
| Z = 4 | 0.15 × 0.12 × 0.04 mm |
| Bruker APEXII CCD diffractometer | 2231 reflections with I > 2σ(I) |
| φ and ω scans | Rint = 0.079 |
| Absorption correction: multi-scan (SADABS; Krause et al., 2015) | θmax = 28.3°, θmin = 2.8° |
| Tmin = 0.415, Tmax = 0.774 | h = −9→9 |
| 97897 measured reflections | k = −10→10 |
| 2562 independent reflections | l = −23→23 |
| Refinement on F2 | Primary atom site location: structure-invariant direct methods |
| Least-squares matrix: full | Hydrogen site location: inferred from neighbouring sites |
| R[F2 > 2σ(F2)] = 0.030 | Only H-atom displacement parameters refined |
| wR(F2) = 0.073 | w = 1/[σ2(Fo2) + (0.0158P)2 + 6.4943P] where P = (Fo2 + 2Fc2)/3 |
| S = 1.28 | (Δ/σ)max = 0.001 |
| 2562 reflections | Δρmax = 1.40 e Å−3 |
| 101 parameters | Δρmin = −0.91 e Å−3 |
| 0 restraints |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
Refinement. The H atoms were refined in idealized geometries and allowed to ride on the parent carbon [d(C—H) = 0.95 Å] and nitrogen atoms [d(N—H) = 0.88 Å] with one common isotropic temperature factor. |
| x | y | z | Uiso*/Ueq | ||
| Sn1 | 0.75694 (5) | 0.43421 (4) | 0.24976 (2) | 0.01293 (9) | |
| Cl1 | 0.78133 (17) | 0.63009 (14) | 0.36546 (6) | 0.0161 (2) | |
| Cl2 | 1.00513 (16) | 0.24850 (15) | 0.32359 (6) | 0.0159 (2) | |
| Cl3 | 0.50030 (16) | 0.27775 (15) | 0.30640 (7) | 0.0180 (2) | |
| Br1 | 0.24791 (7) | 1.05162 (6) | 0.52167 (3) | 0.01847 (12) | |
| N1 | 0.2791 (6) | 0.5686 (5) | 0.3868 (2) | 0.0182 (8) | |
| H1 | 0.2893 | 0.4752 | 0.3616 | 0.034 (8)* | |
| C4 | 0.2545 (6) | 0.8551 (6) | 0.4651 (3) | 0.0137 (9) | |
| C5 | 0.2871 (7) | 0.8616 (6) | 0.3896 (3) | 0.0174 (10) | |
| H5 | 0.2998 | 0.9652 | 0.3648 | 0.034 (8)* | |
| C6 | 0.3003 (7) | 0.7125 (6) | 0.3517 (3) | 0.0177 (10) | |
| H6 | 0.3248 | 0.7128 | 0.3003 | 0.034 (8)* | |
| C2 | 0.2428 (8) | 0.5598 (7) | 0.4590 (3) | 0.0220 (11) | |
| H2 | 0.2260 | 0.4545 | 0.4817 | 0.034 (8)* | |
| C3 | 0.2299 (8) | 0.7035 (6) | 0.5002 (3) | 0.0204 (11) | |
| H3 | 0.2048 | 0.6993 | 0.5515 | 0.034 (8)* |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| Sn1 | 0.01820 (16) | 0.00819 (14) | 0.01255 (14) | 0.00055 (13) | 0.00242 (11) | −0.00069 (12) |
| Cl1 | 0.0255 (6) | 0.0100 (5) | 0.0128 (5) | 0.0000 (4) | 0.0024 (4) | −0.0014 (4) |
| Cl2 | 0.0160 (5) | 0.0147 (5) | 0.0171 (5) | 0.0021 (4) | 0.0020 (4) | −0.0018 (4) |
| Cl3 | 0.0166 (5) | 0.0159 (5) | 0.0223 (6) | −0.0020 (4) | 0.0052 (4) | −0.0052 (4) |
| Br1 | 0.0200 (2) | 0.0136 (2) | 0.0216 (2) | −0.00019 (19) | 0.00136 (18) | −0.00675 (19) |
| N1 | 0.023 (2) | 0.015 (2) | 0.0169 (19) | 0.0013 (18) | 0.0026 (16) | −0.0060 (17) |
| C4 | 0.013 (2) | 0.012 (2) | 0.016 (2) | 0.0024 (17) | 0.0010 (17) | −0.0023 (18) |
| C5 | 0.022 (3) | 0.013 (2) | 0.017 (2) | 0.0022 (19) | 0.0036 (19) | −0.0010 (19) |
| C6 | 0.019 (2) | 0.021 (3) | 0.014 (2) | −0.0008 (19) | 0.0030 (18) | −0.0022 (19) |
| C2 | 0.037 (3) | 0.012 (2) | 0.018 (2) | −0.002 (2) | 0.005 (2) | 0.000 (2) |
| C3 | 0.031 (3) | 0.021 (3) | 0.011 (2) | 0.001 (2) | 0.008 (2) | −0.0009 (19) |
| Sn1—Cl3 | 2.5475 (12) | C4—C5 | 1.388 (7) |
| Sn1—Cl2 | 2.5711 (12) | C5—C6 | 1.381 (7) |
| Sn1—Cl1 | 2.5745 (11) | C5—H5 | 0.9500 |
| Br1—C4 | 1.873 (5) | C6—H6 | 0.9500 |
| N1—C6 | 1.330 (7) | C2—C3 | 1.374 (7) |
| N1—C2 | 1.340 (6) | C2—H2 | 0.9500 |
| N1—H1 | 0.8800 | C3—H3 | 0.9500 |
| C4—C3 | 1.387 (7) | ||
| Cl3—Sn1—Cl2 | 90.73 (4) | C4—C5—H5 | 121.1 |
| Cl3—Sn1—Cl1 | 88.32 (4) | N1—C6—C5 | 120.4 (5) |
| Cl2—Sn1—Cl1 | 88.30 (4) | N1—C6—H6 | 119.8 |
| C6—N1—C2 | 122.7 (4) | C5—C6—H6 | 119.8 |
| C6—N1—H1 | 118.7 | N1—C2—C3 | 119.8 (5) |
| C2—N1—H1 | 118.7 | N1—C2—H2 | 120.1 |
| C3—C4—C5 | 120.8 (4) | C3—C2—H2 | 120.1 |
| C3—C4—Br1 | 119.0 (4) | C2—C3—C4 | 118.5 (4) |
| C5—C4—Br1 | 120.2 (4) | C2—C3—H3 | 120.8 |
| C6—C5—C4 | 117.8 (5) | C4—C3—H3 | 120.8 |
| C6—C5—H5 | 121.1 |
| D—H···A | D—H | H···A | D···A | D—H···A |
| N1—H1···Cl2i | 0.88 | 2.77 | 3.357 (4) | 125 |
| N1—H1···Cl3 | 0.88 | 2.48 | 3.255 (4) | 147 |
| Symmetry code: (i) x−1, y, z. |
| X/Y | d(Sn—X) | d(Sn···Y) | 〈(X—Sn···Y) | Q | BV(Sn···Y) | BV(Sn—X) | p(Sn···Y) |
| Cl1/Cl12 | 2.575 (1) | 3.174 (3) | 167.21 (3)° | 1.23 | 0.05 | 0.60 | 0.49 |
| Cl2/Cl21 | 2.571 (1) | 3.328 (3) | 166.08 (4)° | 1.29 | 0.07 | 0.57 | 0.39 |
| Cl3/Cl31 | 2.548 (1) | 3.479 (3) | 157.11 (4)° | 1.37 | 0.11 | 0.56 | 0.29 |
| Sum | 0.23 | 1.73 |
| Symmetry codes: (1) 3/2 - x, 1/2 - y, 1/2 - z; (2) 3/2 - x, -1/2 - y, 1/2 - z. |
Acknowledgements
We thank the Deutsche Forschungsgemeinschaft and the Government of Lower-Saxony for funding the diffractometer and acknowledge support by Deutsche Forschungsgemeinschaft (DFG) and Open Access Publishing Fund of Osnabrück University.
References
 Adonin, S. A., Gorokh, I. D., Novikov, A. S., Samsonenko, D. G., Yushina, I. V., Sokolov, M. N. & Fedin, V. P. (2018). CrystEngComm 20, 7766–7772. CrossRef CAS Google Scholar
Bauzá, A., Seth, S. K. & Frontera, A. (2019). Coord. Chem. Rev. 384, 107–125. Google Scholar
Brammer, L., Peuronen, A. & Roseveare, T. M. (2023). Acta Cryst. C79, 204–216. Web of Science CrossRef IUCr Journals Google Scholar
Brandenburg, K. (2006). DIAMOND. Crystal Impact GbR, Bonn, Germany. Google Scholar
Brese, N. E. & O'Keeffe, M. (1991). Acta Cryst. B47, 192–197. CrossRef CAS Web of Science IUCr Journals Google Scholar
Bruker (2009). APEX2 and SAINT . Bruker AXS Inc., Madison, Wisconsin, USA. Google Scholar
Cordero, B., Gómez, V., Platero-Prats, A. E., Revés, M., Echeverría, J., Cremades, E., Barragán, F. & Alvarez, S. (2008). Dalton Trans. pp. 2832. Google Scholar
Drew, M. G. B. & Nicholson, D. G. (1986). J. Chem. Soc. Dalton Trans. 15, 1543–1549. CrossRef Google Scholar
Echeverría, J. & Alvarez, S. (2023). Chem. Sci. 14, 11647–11688. Web of Science PubMed Google Scholar
Guthrie, H. D., Stone, S. R. P., Schofield, M. H. & Cahill, C. L. (2026). Cryst. Growth Des. 26, 2325–2338. CrossRef CAS Google Scholar
Izod, K., Clark, E. R., Clegg, W. & Harrington, R. W. (2012). J. Organomet. Chem. 31, 246–255. CrossRef CAS Google Scholar
Krause, L., Herbst-Irmer, R., Sheldrick, G. M. & Stalke, D. (2015). J. Appl. Cryst. 48, 3–10. Web of Science CSD CrossRef ICSD CAS IUCr Journals Google Scholar
Liu, X., Ji, C., Wu, Z., Li, L., Han, S., Wang, Y., Sun, Z. & Luo, J. (2019). Chem. Eur. J. 25, 2610–2615. CrossRef CAS PubMed Google Scholar
Macrae, C. F., Sovago, I., Cottrell, S. J., Galek, P. T. A., McCabe, P., Pidcock, E., Platings, M., Shields, G. P., Stevens, J. S., Towler, M. & Wood, P. A. (2020). J. Appl. Cryst. 53, 226–235. Web of Science CrossRef CAS IUCr Journals Google Scholar
Mantina, M., Chamberlin, A. C., Valero, R., Cramer, C. J. & Truhlar, D. G. (2009). J. Phys. Chem. A 113, 5806–5812. Web of Science CrossRef PubMed CAS Google Scholar
Mínguez Espallargas, G., Brammer, L. & Sherwood, P. (2006). Angew. Chem. Int. Ed. 45, 435–440. Google Scholar
Müller, U., Mronga, N., Schuhmacher, C. & Dehnike, K. (1982). Z. Naturforsch. B37, 1122–1127. Google Scholar
Nicholas, A. D., Garman, L. C., Albano, N. & Cahill, C. L. (2022). Phys. Chem. Chem. Phys. 24, 15305–15320. CrossRef CAS PubMed Google Scholar
Njua, E. Y., Steiner, A. & Stahl, L. (2010). Inorg. Chem. 49, 2163–2172. CrossRef CAS PubMed Google Scholar
Reuter, H. (2025). Molecules 30, 2700. Web of Science CrossRef PubMed Google Scholar
Schröder, S., Röwekamp-Krugley, N., Imwalle, M. & Reuter, H. (2024). Z. Anorg. Allg. Chem. 650, e202400091. Google Scholar
Sheldrick, G. M. (2008). Acta Cryst. A64, 112–122. Web of Science CrossRef CAS IUCr Journals Google Scholar
Sheldrick, G. M. (2015). Acta Cryst. C71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
Veith, M., Jarczyk, M. & Huch, M. (1988). Chem. Ber. 121, 347–355. CrossRef CAS Google Scholar
Walusiak, B. W., Raghavan, A. & Cahill, C. L. (2023). RSC Adv. 13, 13477–13492. CrossRef CAS PubMed Google Scholar
Westrip, S. P. (2010). J. Appl. Cryst. 43, 920–925. Web of Science CrossRef CAS IUCr Journals Google Scholar
Willett, R. D., Awwadi, F., Butcher, R., Haddad, S. & Twamley, B. (2003). Cryst. Growth Des. 3, 301–311. Web of Science CrossRef CAS Google Scholar
Zordan, F., Purver, S. L., Adams, H. & Brammer, L. (2005). CrystEngComm 7, 350–354. CrossRef CAS Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| IUCrDATA |