

 menu


organic compounds
 | IUCrDATA |
 access
1-Naphthamide

aEnvironmental, Molecular and Structural Chemistry Research Unit, University of Constantine-1, 25000, Constantine, Algeria, bLaboratoire de Technologie des Matériaux Avancés, Ecole Nationale, Polytechnique, de Constantine, Algeria, and cCRM2, UMR CNRS 7036, Faculté des Sciences et Technologies, Université de Lorraine, Institut Jean Barriol, BP 70239, 54506 Vandoeuvre-lès-Nancy Cedex, France
*Correspondence e-mail: [email protected]
In the title compound, C11H9NO, the dihedral angle between the naphthalene fused-ring system and the amide group is 42.18 (9)°. In the crystal, N—H⋯O hydrogen bonds link the molecules into (100) sheets. No significant aromatic π–π stacking is observed.
Keywords: crystal structure; naphthalene ring; hydrogen bonds.
CCDC reference: 2590330
Structure description
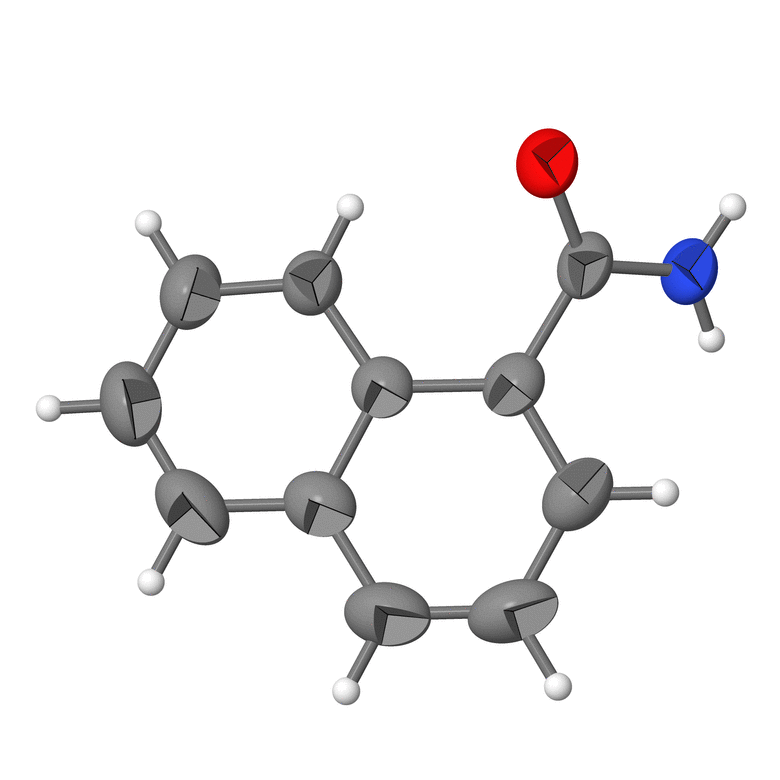
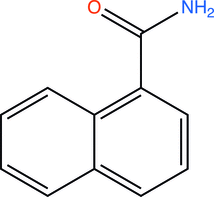
The title compound (I), with molecular formula C11H9NO, crystallizes in the monoclinic space group P21/c. The asymmetric unit contains one molecule in which the dihedral angle between the C2–C11 fused ring system (r.m.s. deviation = 0.008 Å) and the C1/N1/O1 amide moiety is 42.18 (9)° (Fig. 1 ). The significant twist probably arises to minimize steric crowding. The geometrical parameters for the amide group [C1—N1 = 1.3249 (19) Å, C1—O1 = 1.245 (2) Å and angles O1—C1—N1, O1—C1—C2 and N1—C1—C2 close to 120°) support a conjugated amide system, as expected. A short intramolecular C4—H4⋯O1 contact occurs.
 | Figure 1 The molecular structure of (I) with displacement ellipsoids drawn at the 50% probability level. |
In the extended structure of (I), the crystal cohesion is dominated by N11—H1A⋯O1 and N1—H1B⋯O1 hydrogen bonds (Table 1). The former generate inversion dimers and the latter link the dimers into (100) sheets (Fig. 2 and Table 1). The separations between the aromatic ring planes [shortest centroid–centroid separation = 4.799 (4) Å] are too large to support significant aromatic π–π stacking interactions.
| |||||||||||||||||||||||||||
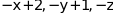 ; (ii)
 .
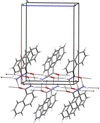 | Figure 2 The packing in (I) with hydrogen bonds shown as black dashed lines. |
Hirshfeld surface analysis and the associated two-dimensional fingerprint plots were used to quantify the intermolecular contacts in the crystal of (I) (Fig. 3). The fingerprint plots show that H⋯H contacts make the largest contribution (42.2%), followed closely by C⋯H/H⋯C contacts (40.2%). The O⋯H/H⋯O contacts account for 14.20%, while all other contributions are less than 2.0%. These results show that the cohesion of the crystal is primarily governed by van der Waals contacts, with aromatic π–π stacking interactions not playing a significant role in establishing the supramolecular architecture.
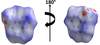 | Figure 3 Hirshfeld surfaces of (I) mapped over dnorm, shown as front and back views. |
For the related structure of 9-anthramide, see Heller & Schmidt (1971).
Synthesis and crystallization
1-Naphthamide was obtained by aqueous hydrolysis of 1-naphthonitrile under heating. The starting nitrile was suspended in water and the mixture was heated until complete conversion to the amide was achieved. Colourless blocks of (I) were grown by slow evaporation of an aqueous solution of the compound at room temperature.
Refinement
Crystal data, data collection and structure details are summarized in Table 2.
|
Structural data
CCDC reference: 2590330
contains datablock I. DOI: https://doi.org/10.1107/S2414314626009892/hb4572sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S2414314626009892/hb4572Isup2.hkl
Supporting information file. DOI: https://doi.org/10.1107/S2414314626009892/hb4572Isup3.cml
| C11H9NO | F(000) = 360 |
| Mr = 171.19 | Dx = 1.276 Mg m−3 |
| Monoclinic, P21/c | Mo Kα radiation, λ = 0.71073 Å |
| a = 14.787 (5) Å | Cell parameters from 2494 reflections |
| b = 6.053 (5) Å | θ = 2.5–35.6° |
| c = 9.954 (5) Å | µ = 0.08 mm−1 |
| β = 90.448 (5)° | T = 293 K |
| V = 890.9 (9) Å3 | Block, colourless |
| Z = 4 | 0.14 × 0.13 × 0.11 mm |
| Saxi-CrysAlisPro-abstract goniometer imported SAXI images diffractometer | 2584 independent reflections |
| Radiation source: fine-focus sealed X-ray tube, Enhance (Mo) X-ray Source | 1535 reflections with I > 2σ(I) |
| Graphite monochromator | Rint = 0.075 |
| ω scans | θmax = 30.0°, θmin = 3.6° |
| Absorption correction: multi-scan (CrysAlisPro; Rigaku OD, 2015) | h = −9→20 |
| Tmin = 0.753, Tmax = 1.000 | k = −8→5 |
| 5594 measured reflections | l = −13→13 |
| Refinement on F2 | Primary atom site location: dual |
| Least-squares matrix: full | Hydrogen site location: inferred from neighbouring sites |
| R[F2 > 2σ(F2)] = 0.071 | H-atom parameters constrained |
| wR(F2) = 0.207 | w = 1/[σ2(Fo2) + (0.106P)2 + 0.0779P] where P = (Fo2 + 2Fc2)/3 |
| S = 1.05 | (Δ/σ)max < 0.001 |
| 2584 reflections | Δρmax = 0.22 e Å−3 |
| 118 parameters | Δρmin = −0.27 e Å−3 |
| 0 restraints |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
Refinement. The H atoms were placed geometrically (N—H = 0.86 Å, C—H = 0.93 Å) and refined as riding atoms. |
| x | y | z | Uiso*/Ueq | ||
| O1 | 0.92484 (9) | 0.3228 (2) | 0.10353 (11) | 0.0589 (4) | |
| C11 | 0.84713 (15) | −0.1053 (3) | −0.10734 (18) | 0.0600 (5) | |
| H11 | 0.9018 | −0.1167 | −0.1524 | 0.072* | |
| C2 | 0.83223 (12) | 0.0721 (3) | −0.02377 (15) | 0.0456 (4) | |
| C1 | 0.90457 (11) | 0.2427 (3) | −0.00788 (15) | 0.0472 (4) | |
| C3 | 0.74867 (11) | 0.0887 (3) | 0.04678 (15) | 0.0439 (4) | |
| C4 | 0.72663 (12) | 0.2652 (3) | 0.13314 (17) | 0.0509 (4) | |
| H4 | 0.7684 | 0.3780 | 0.1470 | 0.061* | |
| C8 | 0.68255 (13) | −0.0809 (3) | 0.02734 (19) | 0.0550 (5) | |
| C7 | 0.59888 (15) | −0.0640 (4) | 0.0951 (2) | 0.0721 (6) | |
| H7 | 0.5555 | −0.1735 | 0.0827 | 0.086* | |
| C9 | 0.70104 (17) | −0.2580 (3) | −0.0600 (2) | 0.0681 (6) | |
| H9 | 0.6579 | −0.3681 | −0.0725 | 0.082* | |
| C5 | 0.64530 (15) | 0.2736 (4) | 0.1967 (2) | 0.0659 (6) | |
| H5 | 0.6326 | 0.3912 | 0.2537 | 0.079* | |
| C10 | 0.78034 (18) | −0.2706 (3) | −0.1257 (2) | 0.0701 (6) | |
| H10 | 0.7912 | −0.3884 | −0.1833 | 0.084* | |
| C6 | 0.58088 (15) | 0.1078 (5) | 0.1774 (2) | 0.0772 (7) | |
| H6 | 0.5256 | 0.1156 | 0.2210 | 0.093* | |
| N1 | 0.94732 (11) | 0.3035 (3) | −0.11836 (13) | 0.0605 (5) | |
| H1A | 0.9900 | 0.3999 | −0.1139 | 0.073* | |
| H1B | 0.9324 | 0.2465 | −0.1945 | 0.073* |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| O1 | 0.0605 (8) | 0.0885 (10) | 0.0276 (6) | −0.0165 (7) | 0.0030 (5) | −0.0033 (6) |
| C11 | 0.0731 (13) | 0.0653 (12) | 0.0416 (9) | 0.0114 (10) | 0.0017 (8) | −0.0031 (8) |
| C2 | 0.0543 (10) | 0.0526 (10) | 0.0299 (7) | 0.0032 (7) | −0.0019 (6) | 0.0028 (7) |
| C1 | 0.0478 (9) | 0.0653 (11) | 0.0285 (7) | 0.0027 (7) | 0.0025 (6) | 0.0008 (7) |
| C3 | 0.0499 (9) | 0.0482 (9) | 0.0335 (7) | −0.0009 (7) | −0.0040 (6) | 0.0066 (6) |
| C4 | 0.0527 (10) | 0.0581 (10) | 0.0418 (9) | −0.0015 (8) | 0.0036 (7) | 0.0006 (7) |
| C8 | 0.0630 (11) | 0.0545 (10) | 0.0472 (9) | −0.0073 (8) | −0.0089 (8) | 0.0095 (8) |
| C7 | 0.0613 (13) | 0.0844 (15) | 0.0703 (14) | −0.0239 (11) | −0.0050 (10) | 0.0150 (12) |
| C9 | 0.0880 (16) | 0.0549 (12) | 0.0612 (13) | −0.0146 (10) | −0.0138 (11) | 0.0033 (9) |
| C5 | 0.0580 (11) | 0.0825 (14) | 0.0572 (12) | 0.0048 (10) | 0.0090 (9) | −0.0071 (10) |
| C10 | 0.1031 (18) | 0.0500 (11) | 0.0572 (12) | 0.0029 (11) | −0.0113 (12) | −0.0088 (9) |
| C6 | 0.0538 (12) | 0.1050 (18) | 0.0730 (15) | −0.0059 (12) | 0.0124 (10) | 0.0030 (13) |
| N1 | 0.0655 (10) | 0.0879 (11) | 0.0281 (7) | −0.0179 (8) | 0.0101 (6) | −0.0073 (7) |
| O1—C1 | 1.245 (2) | C8—C9 | 1.408 (3) |
| C11—H11 | 0.9300 | C7—H7 | 0.9300 |
| C11—C2 | 1.377 (3) | C7—C6 | 1.352 (3) |
| C11—C10 | 1.417 (3) | C9—H9 | 0.9300 |
| C2—C1 | 1.495 (3) | C9—C10 | 1.349 (3) |
| C2—C3 | 1.430 (2) | C5—H5 | 0.9300 |
| C1—N1 | 1.3249 (19) | C5—C6 | 1.396 (3) |
| C3—C4 | 1.411 (2) | C10—H10 | 0.9300 |
| C3—C8 | 1.430 (3) | C6—H6 | 0.9300 |
| C4—H4 | 0.9300 | N1—H1A | 0.8600 |
| C4—C5 | 1.364 (3) | N1—H1B | 0.8600 |
| C8—C7 | 1.417 (3) | ||
| C2—C11—H11 | 119.5 | C8—C7—H7 | 119.3 |
| C2—C11—C10 | 120.9 (2) | C6—C7—C8 | 121.37 (19) |
| C10—C11—H11 | 119.5 | C6—C7—H7 | 119.3 |
| C11—C2—C1 | 119.01 (17) | C8—C9—H9 | 119.5 |
| C11—C2—C3 | 119.67 (17) | C10—C9—C8 | 121.05 (19) |
| C3—C2—C1 | 121.32 (15) | C10—C9—H9 | 119.5 |
| O1—C1—C2 | 122.04 (14) | C4—C5—H5 | 119.6 |
| O1—C1—N1 | 121.23 (17) | C4—C5—C6 | 120.8 (2) |
| N1—C1—C2 | 116.73 (14) | C6—C5—H5 | 119.6 |
| C2—C3—C8 | 118.39 (16) | C11—C10—H10 | 119.8 |
| C4—C3—C2 | 123.88 (15) | C9—C10—C11 | 120.35 (19) |
| C4—C3—C8 | 117.72 (17) | C9—C10—H10 | 119.8 |
| C3—C4—H4 | 119.3 | C7—C6—C5 | 120.0 (2) |
| C5—C4—C3 | 121.30 (18) | C7—C6—H6 | 120.0 |
| C5—C4—H4 | 119.3 | C5—C6—H6 | 120.0 |
| C7—C8—C3 | 118.83 (18) | C1—N1—H1A | 120.0 |
| C9—C8—C3 | 119.63 (19) | C1—N1—H1B | 120.0 |
| C9—C8—C7 | 121.54 (19) | H1A—N1—H1B | 120.0 |
| C11—C2—C1—O1 | −137.42 (18) | C3—C8—C7—C6 | −0.2 (3) |
| C11—C2—C1—N1 | 41.7 (2) | C3—C8—C9—C10 | 0.1 (3) |
| C11—C2—C3—C4 | −179.15 (16) | C4—C3—C8—C7 | 0.0 (2) |
| C11—C2—C3—C8 | −0.3 (2) | C4—C3—C8—C9 | 178.83 (16) |
| C2—C11—C10—C9 | −0.7 (3) | C4—C5—C6—C7 | 0.2 (4) |
| C2—C3—C4—C5 | 179.25 (17) | C8—C3—C4—C5 | 0.4 (3) |
| C2—C3—C8—C7 | −178.98 (16) | C8—C7—C6—C5 | 0.1 (4) |
| C2—C3—C8—C9 | −0.1 (2) | C8—C9—C10—C11 | 0.3 (3) |
| C1—C2—C3—C4 | 1.4 (2) | C7—C8—C9—C10 | 178.9 (2) |
| C1—C2—C3—C8 | −179.74 (15) | C9—C8—C7—C6 | −179.1 (2) |
| C3—C2—C1—O1 | 42.0 (2) | C10—C11—C2—C1 | −179.83 (17) |
| C3—C2—C1—N1 | −138.82 (17) | C10—C11—C2—C3 | 0.7 (3) |
| C3—C4—C5—C6 | −0.5 (3) |
| D—H···A | D—H | H···A | D···A | D—H···A |
| C4—H4···O1 | 0.93 | 2.38 | 2.969 (3) | 121 |
| N1—H1A···O1i | 0.86 | 2.10 | 2.951 (3) | 170 |
| N1—H1B···O1ii | 0.86 | 2.06 | 2.889 (3) | 163 |
| Symmetry codes: (i) −x+2, −y+1, −z; (ii) x, −y+1/2, z−1/2. |
Acknowledgements
The authors thank the Cambridge Crystallographic Data Center (CCDC) for access to the Cambridge Structural Database (CSD) through the FAIRE program.
Funding information
Funding for this research was provided by: Unité de recherche de chimie de l'environnement, moléculaire et structurale UR.CHEMS; DRSDT-Algeria; Laboratoire de technologie des matériaux avancés, Ecole Nationale Polytechnique de Constantine, Algeria.
References
 Bourhis, L. J., Dolomanov, O. V., Gildea, R. J., Howard, J. A. K. & Puschmann, H. (2015). Acta Cryst. A71, 59–75. Web of Science CrossRef IUCr Journals Google Scholar
Dolomanov, O. V., Bourhis, L. J., Gildea, R. J., Howard, J. A. K. & Puschmann, H. (2009). J. Appl. Cryst. 42, 339–341. Web of Science CrossRef CAS IUCr Journals Google Scholar
Heller, E. & Schmidt, G. M. J. (1971). Isr. J. Chem. 9, 449–462. CSD CrossRef CAS Google Scholar
Rigaku OD (2015). CrysAlis PRO. Rigaku Oxford Diffraction, Yarnton, England. Google Scholar
Sheldrick, G. M. (2015). Acta Cryst. C71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| IUCrDATA |