

 menu


metal-organic compounds
 | IUCrDATA |
 access
Bis(pyridinium) tetrachloridozincate(II), [Hpy]2[ZnCl4]

aChemistry, Osnabrück University, Barbarastr. 7, 49069 Osnabrück, Germany
*Correspondence e-mail: [email protected]
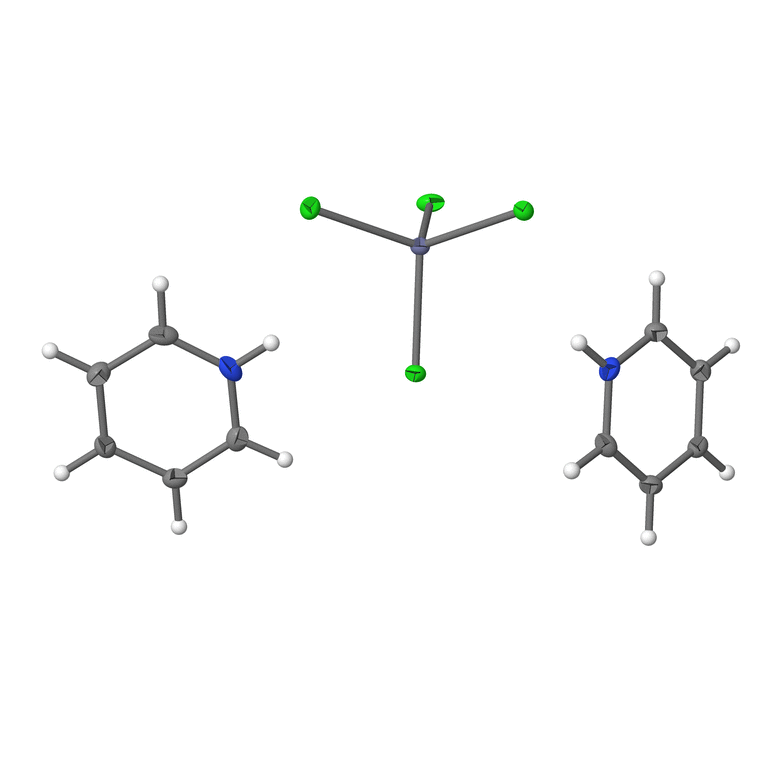
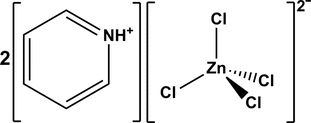
Single crystals of the title compound, (C5H6N)2[ZnCl4] or [Hpy]2[ZnCl4], were grown from an aqueous solution of pyridinium chloride and zinc dichloride. The compound is isostructural with the corresponding cobalt compound and exhibits an almost tetrahedral [ZnCl4]2− anion and two different pyridinium cations. The structure is described with respect to the bond lengths and angles of the anion and the π–π interactions between the cations, whilst mention is made of the N—H⋯Cl and C—H⋯Cl interactions already known from the cobalt compound.
CCDC reference: 2590726
Structure description
The title compound is isostructural with the analogous cobalt compound (Felloni et al., 2004 ) and crystallizes in the triclinic space group P
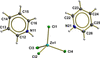
 with one formula unit in the asymmetric unit (Fig. 1). The zinc atom is almost tetrahedrally [〈(Cl—Zn—Cl) = 105.62 (1)–111.91 (1)°, 〈(Cl—Zn—Cl)mean = 109.5 (23)°] coordinated from four chlorine atoms. The zinc–chlorine distances are, on average, slightly shorter [d(Zn—Cl)mean = 2.273 (11) Å] than the cobalt-chlorine distances [d(Co—Cl)mean = 2.276 (7) Å], but vary much more widely [2.2605 (3) − 2.2862 (3) Å] than the latter. Zn—Cl distances of similar length and variation can be found, for example, in the compound with the 4,4′-bipyridinium ion [d(Zn—Cl)mean = 2.275 (27) Å] (Gillon et al., 2000) and in the compound with the 3-chloropyridinium ion [d(Zn—Cl)mean = 2.272 (14) Å] (Mínguez Espallargas et al., 2009).
|
Figure 1
Asymmetric unit of the crystal structure of [Hpy]2[ZnCl4] with atom numbering according to the isostructural cobalt compound. With the exception of the hydrogen atoms, which are shown as spheres of arbitrary radius, all other atoms are drawn as anisotropic displacement ellipsoids at the 50% probability level. |
As the publication on the cobalt compound (Felloni et al., 2004) describes the key aspects of the crystal structure, particularly with regard to the pattern of N—H⋯Cl hydrogen bonds and C—H⋯Cl contacts, there is no need to provide a detailed description of this here. All the necessary information to these supramolecular interactions can be found in Table 1 and can be compared directly with the values reported for the cobalt compound, as the asymmetric unit has been adjusted in terms of atomic numbering and its position within the unit cell.
|
 ; (ii)
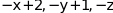 ; (iii)
 ; (iv)
 ; (v)
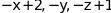 .
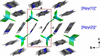
However, another aspect is presented here that is not given sufficient attention in the publication on the cobalt compound. Within the crystal structure (Fig. 2), the two pyridinium ions are arranged such that they partially overlap: Hpy(1) on one side, Hpy(2) on both sides. In all three cases, the relevant element of symmetry is a center of symmetry. In the case of Hpy(1), this is located at Wyckhoff position e; in the case of Hpy(2), at Wyckhoff positions c and h. The centers of symmetry ensure that the planes of the two neighboring pyridinium ions are exactly parallel to one another, with the nitrogen atoms facing in opposite directions.
|
Figure 2
Ball-and-stick model of the ions in the crystal structure of the title compound showing their three-dimensional packing when looking down the a axis. The layers of the two different pyridinium ions are labeled and the relevant centers of symmetry (black dots) marked with Wyckhoff letters. Color code: Zn = light blue, Cl = yellow–green, N = dark blue, C = dark gray, H = white. |
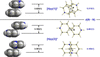
It can be seen from Fig. 3 that the more the pyridinium ions overlap, the greater the distance between the planes becomes. The different distances between the nitrogen atoms in these arrangements is also evident.
|
Figure 3
Different representations of the cation stacking. (left) Space-filling model with stick model overlayed; side-view with distances between the least-squares planes (Å, black values). (right) Ball-and-stick models of the centrosymmetric dimers looking down on the least-square planes with distances (Å, blue values])between the nitrogen atoms and positions of the relevant centers of symmetry (black dots) marked with the Wyckhoff letters. |
Synthesis and crystallization
The title compound was synthesized by dissolving equimolar quantities of zinc chloride and pyridinium chloride in water. When the solvent evaporates, the compound crystallizes in transparent, slightly green needles.
Refinement
Crystal data,data collection and structure refinement details are summarized in Table 2. Once it had become apparent that the compound was isostructural with the cobalt compound, its asymmetric unit was used to describe the structure.
|
Structural data
CCDC reference: 2590726
Crystal structure: contains datablock I. DOI: https://doi.org/10.1107/S241431462601000X/bv4063sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S241431462601000X/bv4063Isup2.hkl
| (C5H6N)2[ZnCl4] | Z = 2 |
| Mr = 367.39 | F(000) = 368 |
| Triclinic, P1 | Dx = 1.675 Mg m−3 |
| a = 7.5711 (3) Å | Mo Kα radiation, λ = 0.71073 Å |
| b = 7.9451 (3) Å | Cell parameters from 9982 reflections |
| c = 12.3798 (5) Å | θ = 2.6–29.6° |
| α = 88.942 (2)° | µ = 2.40 mm−1 |
| β = 83.518 (2)° | T = 100 K |
| γ = 79.959 (2)° | Needle, green |
| V = 728.59 (5) Å3 | 0.39 × 0.24 × 0.19 mm |
| Bruker APEXII CCD diffractometer | 3846 reflections with I > 2σ(I) |
| φ and ω scans | Rint = 0.030 |
| Absorption correction: multi-scan (SADABS; Krause et al., 2015) | θmax = 29.6°, θmin = 2.6° |
| Tmin = 0.498, Tmax = 0.523 | h = −10→10 |
| 82340 measured reflections | k = −11→11 |
| 4103 independent reflections | l = −17→17 |
| Refinement on F2 | Primary atom site location: structure-invariant direct methods |
| Least-squares matrix: full | Hydrogen site location: inferred from neighbouring sites |
| R[F2 > 2σ(F2)] = 0.016 | Only H-atom displacement parameters refined |
| wR(F2) = 0.041 | w = 1/[σ2(Fo2) + (0.0205P)2 + 0.2351P] where P = (Fo2 + 2Fc2)/3 |
| S = 1.07 | (Δ/σ)max = 0.002 |
| 4103 reflections | Δρmax = 0.38 e Å−3 |
| 156 parameters | Δρmin = −0.28 e Å−3 |
| 0 restraints |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
Refinement. The H atoms were refined in idealized geometries and allowed to ride on the parent carbon [d(C—H) = 0.95 Å] and nitrogen atoms [d(N—H) = 0.88 Å] with one common isotropic temperature factor for each of the two pyridinium ions. |
| x | y | z | Uiso*/Ueq | ||
| Zn1 | 0.94436 (2) | 0.17307 (2) | 0.22056 (2) | 0.01280 (4) | |
| Cl1 | 0.67207 (3) | 0.09965 (3) | 0.19775 (2) | 0.01488 (5) | |
| Cl2 | 1.10183 (3) | 0.18295 (3) | 0.05356 (2) | 0.01455 (5) | |
| Cl3 | 0.90410 (3) | 0.43146 (3) | 0.30190 (2) | 0.01870 (6) | |
| Cl4 | 1.08368 (3) | −0.03785 (3) | 0.32370 (2) | 0.01898 (6) | |
| N11 | 0.75046 (12) | 0.23776 (12) | −0.07461 (8) | 0.01706 (18) | |
| H11 | 0.8367 | 0.1659 | −0.0479 | 0.0247 (15)* | |
| C12 | 0.77721 (15) | 0.39734 (15) | −0.09665 (9) | 0.0185 (2) | |
| H12 | 0.8877 | 0.4310 | −0.0845 | 0.0247 (15)* | |
| C13 | 0.64300 (15) | 0.51229 (14) | −0.13719 (9) | 0.0185 (2) | |
| H13 | 0.6597 | 0.6262 | −0.1534 | 0.0247 (15)* | |
| C14 | 0.48320 (15) | 0.45976 (14) | −0.15402 (9) | 0.0170 (2) | |
| H14 | 0.3889 | 0.5381 | −0.1812 | 0.0247 (15)* | |
| C15 | 0.46079 (14) | 0.29270 (14) | −0.13117 (9) | 0.0170 (2) | |
| H15 | 0.3518 | 0.2555 | −0.1429 | 0.0247 (15)* | |
| C16 | 0.59836 (15) | 0.18200 (14) | −0.09133 (9) | 0.0181 (2) | |
| H16 | 0.5860 | 0.0668 | −0.0757 | 0.0247 (15)* | |
| N21 | 0.70751 (13) | −0.16465 (12) | 0.40981 (8) | 0.01933 (19) | |
| H21 | 0.7710 | −0.1133 | 0.3597 | 0.0294 (17)* | |
| C22 | 0.53087 (16) | −0.15652 (14) | 0.40250 (9) | 0.0194 (2) | |
| H22 | 0.4764 | −0.0965 | 0.3443 | 0.0294 (17)* | |
| C23 | 0.42975 (15) | −0.23567 (14) | 0.47987 (9) | 0.0188 (2) | |
| H23 | 0.3045 | −0.2314 | 0.4757 | 0.0294 (17)* | |
| C24 | 0.51212 (15) | −0.32184 (14) | 0.56405 (9) | 0.0181 (2) | |
| H24 | 0.4433 | −0.3767 | 0.6183 | 0.0294 (17)* | |
| C25 | 0.69495 (16) | −0.32800 (15) | 0.56911 (9) | 0.0210 (2) | |
| H25 | 0.7528 | −0.3871 | 0.6265 | 0.0294 (17)* | |
| C26 | 0.79136 (15) | −0.24706 (15) | 0.48961 (10) | 0.0211 (2) | |
| H26 | 0.9169 | −0.2498 | 0.4915 | 0.0294 (17)* |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| Zn1 | 0.01176 (6) | 0.01293 (6) | 0.01405 (6) | −0.00272 (4) | −0.00215 (4) | 0.00042 (4) |
| Cl1 | 0.01383 (11) | 0.01502 (11) | 0.01738 (11) | −0.00575 (8) | −0.00378 (9) | 0.00149 (8) |
| Cl2 | 0.01337 (11) | 0.01618 (11) | 0.01457 (11) | −0.00401 (8) | −0.00148 (8) | 0.00135 (8) |
| Cl3 | 0.01431 (11) | 0.01759 (12) | 0.02475 (13) | −0.00271 (9) | −0.00354 (9) | −0.00729 (10) |
| Cl4 | 0.01502 (11) | 0.02239 (13) | 0.01953 (12) | −0.00225 (9) | −0.00462 (9) | 0.00743 (10) |
| N11 | 0.0159 (4) | 0.0177 (4) | 0.0164 (4) | 0.0011 (3) | −0.0035 (3) | 0.0023 (3) |
| C12 | 0.0170 (5) | 0.0223 (5) | 0.0180 (5) | −0.0073 (4) | −0.0037 (4) | 0.0005 (4) |
| C13 | 0.0224 (5) | 0.0154 (5) | 0.0194 (5) | −0.0064 (4) | −0.0049 (4) | 0.0030 (4) |
| C14 | 0.0173 (5) | 0.0170 (5) | 0.0165 (5) | −0.0013 (4) | −0.0042 (4) | 0.0013 (4) |
| C15 | 0.0154 (5) | 0.0191 (5) | 0.0174 (5) | −0.0059 (4) | −0.0014 (4) | −0.0018 (4) |
| C16 | 0.0205 (5) | 0.0141 (5) | 0.0195 (5) | −0.0047 (4) | 0.0006 (4) | 0.0010 (4) |
| N21 | 0.0232 (5) | 0.0171 (4) | 0.0178 (4) | −0.0080 (4) | 0.0051 (4) | −0.0022 (3) |
| C22 | 0.0252 (6) | 0.0161 (5) | 0.0174 (5) | −0.0035 (4) | −0.0043 (4) | 0.0000 (4) |
| C23 | 0.0155 (5) | 0.0189 (5) | 0.0226 (5) | −0.0039 (4) | −0.0036 (4) | −0.0007 (4) |
| C24 | 0.0195 (5) | 0.0186 (5) | 0.0163 (5) | −0.0052 (4) | 0.0007 (4) | −0.0002 (4) |
| C25 | 0.0207 (5) | 0.0246 (6) | 0.0180 (5) | −0.0024 (4) | −0.0054 (4) | 0.0000 (4) |
| C26 | 0.0146 (5) | 0.0231 (6) | 0.0259 (6) | −0.0047 (4) | −0.0007 (4) | −0.0061 (4) |
| Zn1—Cl3 | 2.2605 (3) | C15—H15 | 0.9500 |
| Zn1—Cl4 | 2.2715 (3) | C16—H16 | 0.9500 |
| Zn1—Cl2 | 2.2742 (3) | N21—C26 | 1.3344 (16) |
| Zn1—Cl1 | 2.2862 (3) | N21—C22 | 1.3408 (15) |
| N11—C12 | 1.3369 (15) | N21—H21 | 0.8800 |
| N11—C16 | 1.3418 (14) | C22—C23 | 1.3708 (16) |
| N11—H11 | 0.8800 | C22—H22 | 0.9500 |
| C12—C13 | 1.3759 (16) | C23—C24 | 1.3840 (16) |
| C12—H12 | 0.9500 | C23—H23 | 0.9500 |
| C13—C14 | 1.3847 (15) | C24—C25 | 1.3850 (16) |
| C13—H13 | 0.9500 | C24—H24 | 0.9500 |
| C14—C15 | 1.3873 (15) | C25—C26 | 1.3759 (17) |
| C14—H14 | 0.9500 | C25—H25 | 0.9500 |
| C15—C16 | 1.3719 (15) | C26—H26 | 0.9500 |
| Cl3—Zn1—Cl4 | 111.911 (12) | N11—C16—C15 | 119.44 (10) |
| Cl3—Zn1—Cl2 | 109.683 (11) | N11—C16—H16 | 120.3 |
| Cl4—Zn1—Cl2 | 111.059 (11) | C15—C16—H16 | 120.3 |
| Cl3—Zn1—Cl1 | 110.461 (10) | C26—N21—C22 | 122.99 (10) |
| Cl4—Zn1—Cl1 | 105.621 (11) | C26—N21—H21 | 118.5 |
| Cl2—Zn1—Cl1 | 107.971 (10) | C22—N21—H21 | 118.5 |
| C12—N11—C16 | 123.13 (10) | N21—C22—C23 | 119.32 (10) |
| C12—N11—H11 | 118.4 | N21—C22—H22 | 120.3 |
| C16—N11—H11 | 118.4 | C23—C22—H22 | 120.3 |
| N11—C12—C13 | 119.30 (10) | C22—C23—C24 | 119.27 (10) |
| N11—C12—H12 | 120.4 | C22—C23—H23 | 120.4 |
| C13—C12—H12 | 120.4 | C24—C23—H23 | 120.4 |
| C12—C13—C14 | 119.11 (10) | C23—C24—C25 | 119.95 (10) |
| C12—C13—H13 | 120.4 | C23—C24—H24 | 120.0 |
| C14—C13—H13 | 120.4 | C25—C24—H24 | 120.0 |
| C13—C14—C15 | 120.01 (10) | C26—C25—C24 | 118.84 (11) |
| C13—C14—H14 | 120.0 | C26—C25—H25 | 120.6 |
| C15—C14—H14 | 120.0 | C24—C25—H25 | 120.6 |
| C16—C15—C14 | 118.99 (10) | N21—C26—C25 | 119.63 (10) |
| C16—C15—H15 | 120.5 | N21—C26—H26 | 120.2 |
| C14—C15—H15 | 120.5 | C25—C26—H26 | 120.2 |
| D—H···A | D—H | H···A | D···A | D—H···A |
| N11—H11···Cl2 | 0.88 | 2.51 | 3.211 (1) | 137 |
| N11—H11···Cl2i | 0.88 | 2.73 | 3.350 (1) | 129 |
| C12—H12···Cl2 | 0.95 | 2.97 | 3.443 (1) | 112 |
| C13—H13···Cl2ii | 0.95 | 2.94 | 3.584 (1) | 127 |
| C15—H15···Cl1iii | 0.95 | 2.96 | 3.576 (1) | 124 |
| C16—H16···Cl2i | 0.95 | 2.84 | 3.417 (1) | 120 |
| N21—H21···Cl1 | 0.88 | 2.69 | 3.337 (1) | 132 |
| N21—H21···Cl4 | 0.88 | 2.53 | 3.246 (1) | 139 |
| C22—H22···Cl1 | 0.95 | 2.83 | 3.416 (1) | 121 |
| C22—H22···Cl4iv | 0.95 | 2.97 | 3.592 (1) | 125 |
| C23—H23···Cl4iv | 0.95 | 2.90 | 3.559 (1) | 128 |
| C25—H25···Cl3v | 0.95 | 2.80 | 3.552 (1) | 137 |
| C26—H26···Cl4 | 0.95 | 2.96 | 3.466 (2) | 115 |
| Symmetry codes: (i) −x+2, −y, −z; (ii) −x+2, −y+1, −z; (iii) −x+1, −y, −z; (iv) x−1, y, z; (v) −x+2, −y, −z+1. |
Acknowledgements
The Deutsche Forschungsgemeinschaft and the Government of Lower-Saxony are thanked for the funding of the diffractometer.
References
 Brandenburg, K. (2006). DIAMOND. Crystal Impact GbR, Bonn, Germany.
Google Scholar
Bruker (2009). APEX2 and SAINT. Bruker AXS Inc., Madison, Wisconsin, USA.
Google Scholar
Felloni, M., Hubberstey, P., Wilson, C. & Schröder, M. (2004). CrystEngComm 6, 87–95.
CrossRef
CAS
Google Scholar
Gillon, A. L., Lewis, G. R., Orpen, A. G., Rotter, S., Starbuck, J., Wang, X., Rodríguez-Martín, Y. & Ruiz-Pérez, C. (2000). J. Chem. Soc. Dalton Trans. pp. 3897–3905.
Web of Science
CrossRef
Google Scholar
Krause, L., Herbst-Irmer, R., Sheldrick, G. M. & Stalke, D. (2015). J. Appl. Cryst. 48, 3–10.
Web of Science
CSD
CrossRef
ICSD
CAS
IUCr Journals
Google Scholar
Macrae, C. F., Sovago, I., Cottrell, S. J., Galek, P. T. A., McCabe, P., Pidcock, E., Platings, M., Shields, G. P., Stevens, J. S., Towler, M. & Wood, P. A. (2020). J. Appl. Cryst. 53, 226–235.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Mínguez Espallargas, G., Zordan, F., Arroyo Marín, L., Adams, H., Shankland, K., van de Streek, J. & Brammer, L. (2009). Chem. Eur. J. 15, 7554–7568.
Web of Science
PubMed
Google Scholar
Sheldrick, G. M. (2008). Acta Cryst. A64, 112–122.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Sheldrick, G. M. (2015). Acta Cryst. C71, 3–8.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Westrip, S. P. (2010). J. Appl. Cryst. 43, 920–925.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| IUCrDATA |